ISISNeutronMuon / ISISNeutronMuon/MDMC
Lennard-Jones parameter definitions in interaction_functions.py
- Dominant language
- Python
- Stars
- 4
- Forks
- 0
- Avg merge
- 2d 4h
- Merged PRs (30d)
- 5
Description
**Description of the error**
While trying to add more force field for more complex sugar-molecules I discovered a problem.
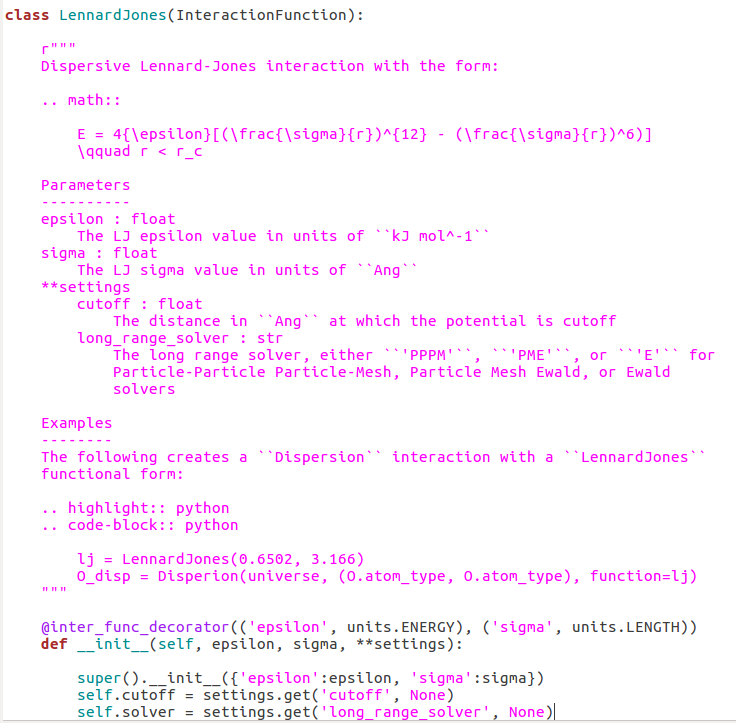
We don't have clear definitions for Lennard-Jones potential values. In the code interaction_functions.py we have defined values or parameter set-ups for charges, angles, dihedrals, which can distinguish more or less between parameters for different atoms, but this option doesn't appear clear for Lennard-Jones potential. I general it looks like we can have only one value for the whole simulation which we can respectively give for one atom.
**Describe the expected result**
We supposed to have options for different values for different atom-pairs
**Describe the actual result**
In the actual result we input only one value.
**Suggested fix**
Make it for multiple values and well-distinguishable.
**Additional details**
The piece of code is here on the image
Contributor guide
No contributing guide indexed for this repository
Assessment
This issue has not been assessed yet.